Medical Device Regulatory Due Diligence: A Market-Entry Document Checklist for Buyers
FDA 510(k) Clearance: What U.S. Buyers Should Know About INTCO Medical Devices
FROM: INTCO Medical|24 Oct 2026
For U.S. buyers evaluating INTCO Medical devices, FDA 510(k) clearance is an important regulatory consideration. Which products require 510(k) clearance, which may be exempt, and what documentation should buyers review before purchasing?
INTCO Medical’s medical device portfolio includes examination gloves, rehabilitation equipment, medical consumables, and cold and hot therapy products, each subject to applicable U.S. regulatory requirements.However, an FDA mark does not establish 510(k) clearance or FDA approval. The applicable U.S. regulatory pathway may involve establishment registration and device listing, 510(k) clearance, or a 510(k) exemption, depending on the product. Buyers should verify the exact SKU, intended use, product code, legal manufacturer, current labeling, and applicable FDA records before making regulatory claims.
What Does FDA 510(k) Clearance Mean for a U.S. Buyer?
FDA 510(k) clearance comes from the 510(k) premarket notification pathway. The submission compares a new device with a legally marketed predicate and supports substantial equivalence for a defined intended use. That decision belongs to a specific device, applicant, and set of indications for use. It does not automatically cover every product in a supplier’s catalogue.
FDA Clearance, Registration, and Listing
FDA establishment registration and device listing identify registered establishments and listed devices, but they do not constitute FDA clearance or approval. A 510(k) clearance is a separate premarket determination of substantial equivalence. Certain devices may be exempt from 510(k) requirements, subject to applicable classification regulations and exemption limitations. Buyers should distinguish registration, listing, clearance, and exemption when reviewing supplier documentation.
Record | What you should confirm |
510(k) clearance | K number, applicant, device identification, indications for use, decision date, and product code, when clearance is applicable |
Establishment registration | Registered establishment, registration status, and establishment owner or operator |
Device listing | Listed device name, product code, associated establishment, and available device identification details |
Catalogue FDA-related mark | Verify the applicable registration, listing, clearance, or exemption evidence. An FDA-related mark alone does not establish FDA approval or clearance. |
Why the Exact SKU Matters
Different medical device models within the same product category may have different materials, intended uses, regulatory classifications, and labeling requirements. Ask for the commercial SKU, size, packaging, intended use, indications for use, and target channel. A family name alone leaves too much room for a mismatch.
FDA Regulatory Status Across INTCO Medical Product Categories
INTCO Medical supplies multiple categories of medical products to international markets, including medical examination gloves, rehabilitation equipment, protective medical consumables, medical electrodes, and cold and hot therapy products. For U.S. buyers, the applicable FDA regulatory pathway depends on the specific device classification, intended use, and product configuration.
FDA 510(k) clearance, establishment registration and device listing, and 510(k) exemption are different regulatory concepts. INTCO Medical’s product portfolio includes devices associated with these different pathways. Buyers should verify the applicable FDA records and supporting documentation for each specific product rather than assuming that a single clearance or registration covers an entire product category.
Medical Examination Gloves
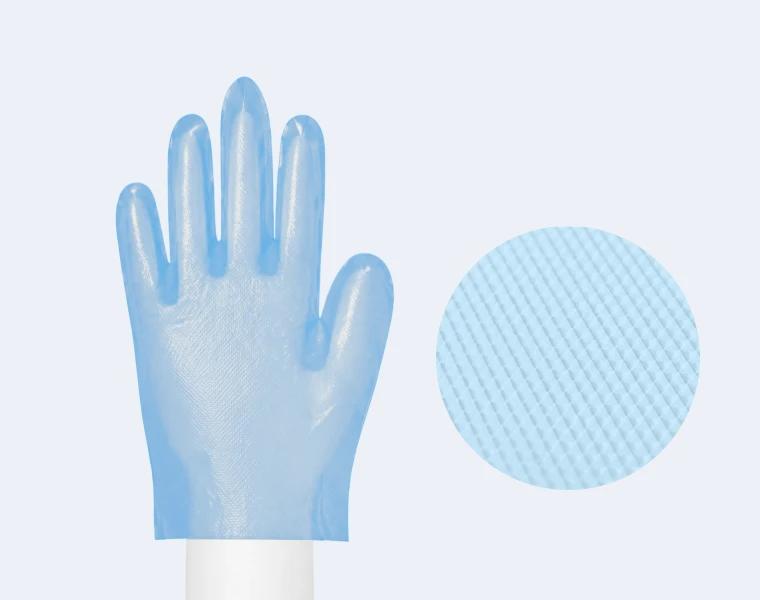
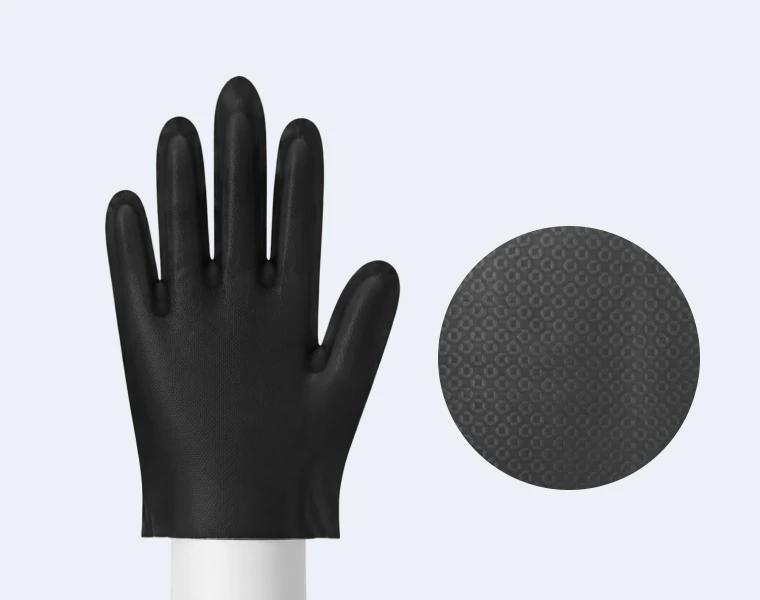
Medical examination gloves are an important part of INTCO Medical’s product portfolio. The company supplies disposable examination gloves, including nitrile and vinyl (PVC) examination gloves, for healthcare and professional applications.
FDA 510(k) records have been identified for selected medical examination gloves within INTCO Medical’s product portfolio. For example, FDA record K231408 identifies a nitrile examination glove submission associated with the product name SynGuard and product code LZA. Additional FDA 510(k) records have been identified for selected nitrile and vinyl examination gloves. The applicability of each clearance should be verified against the specific product and legal manufacturer.
Rehabilitation and Mobility Equipment
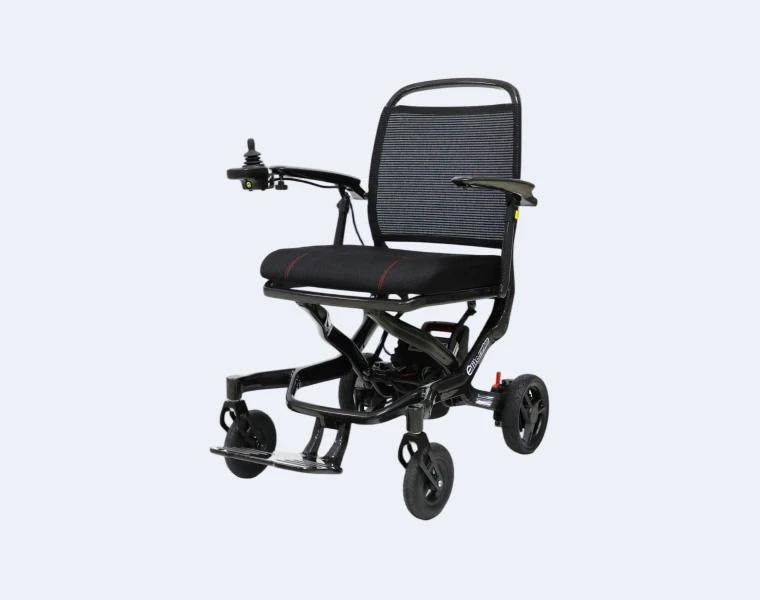
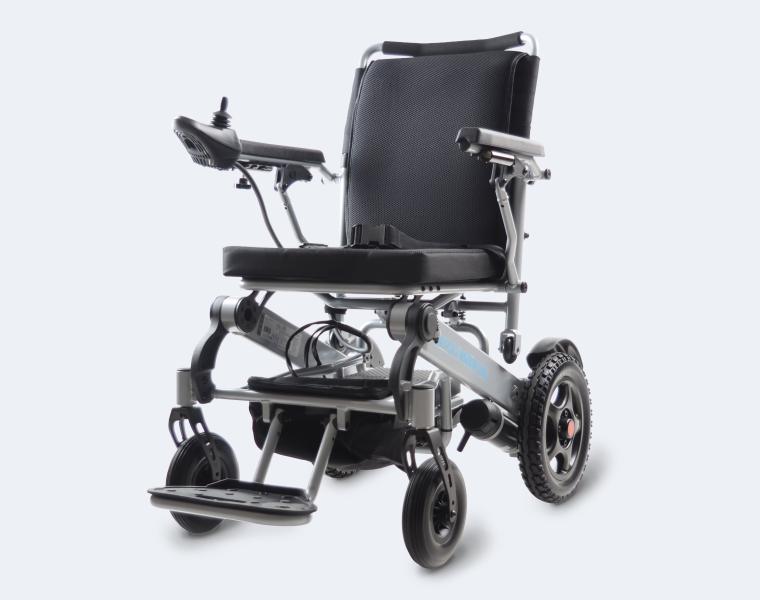
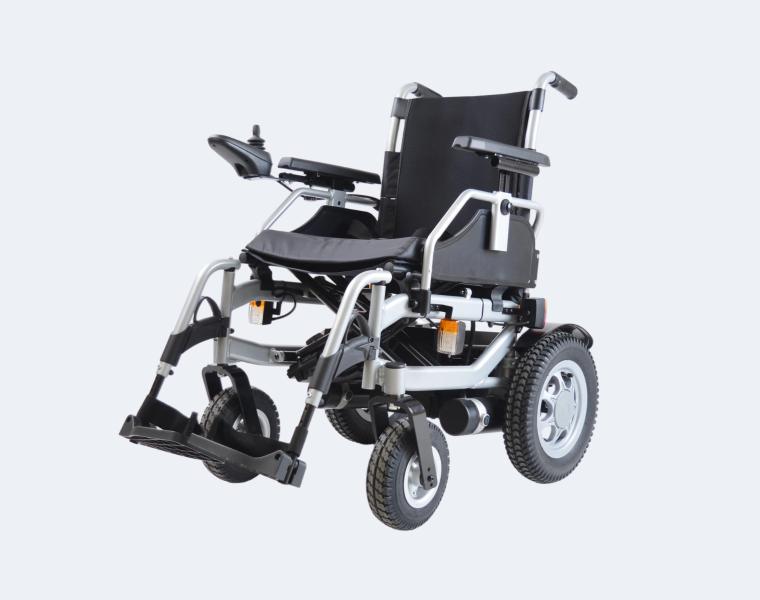
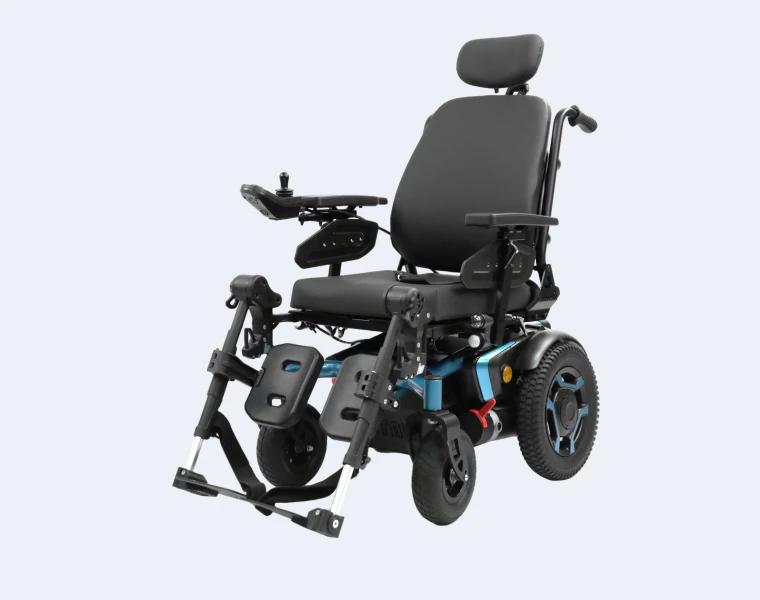
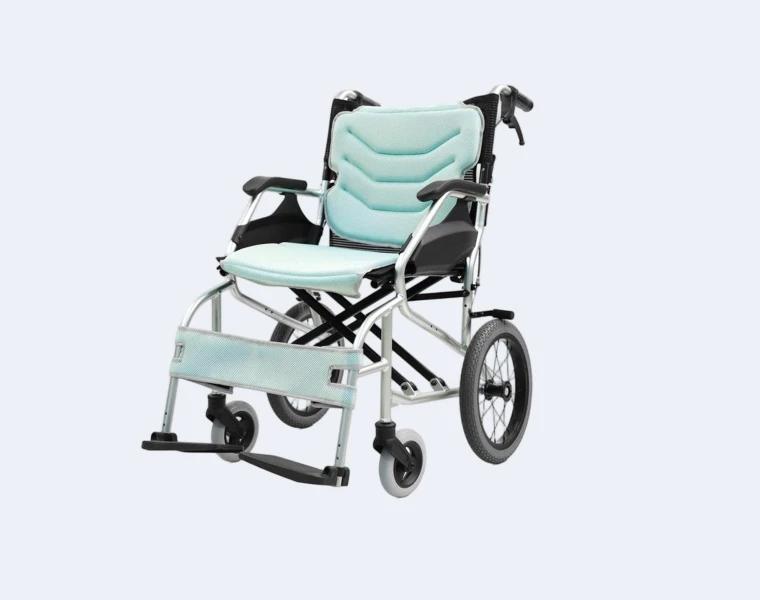
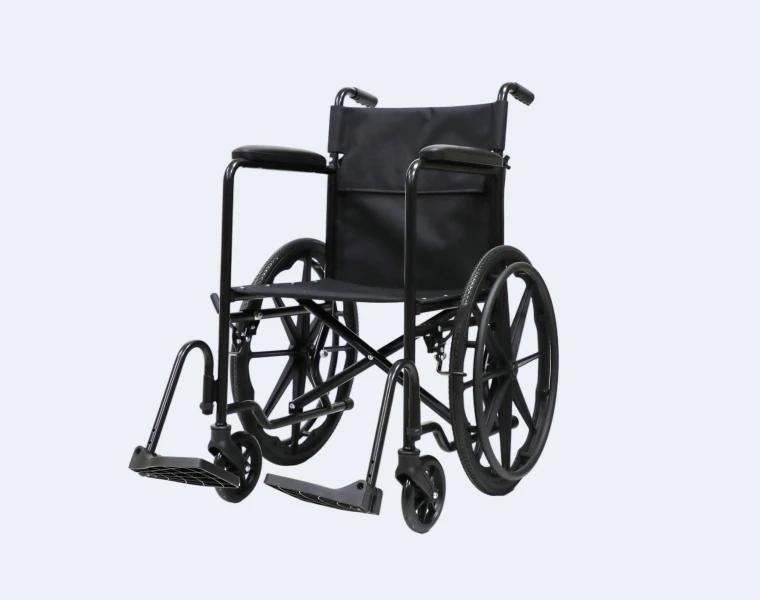
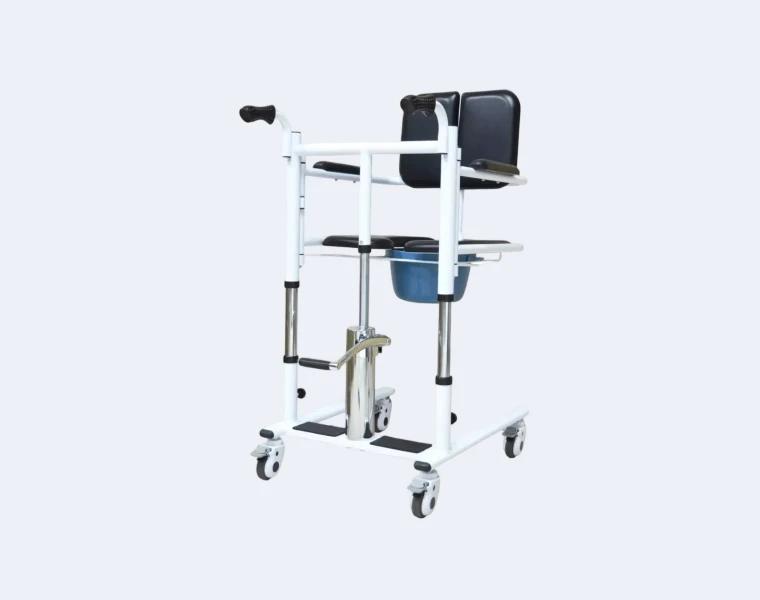
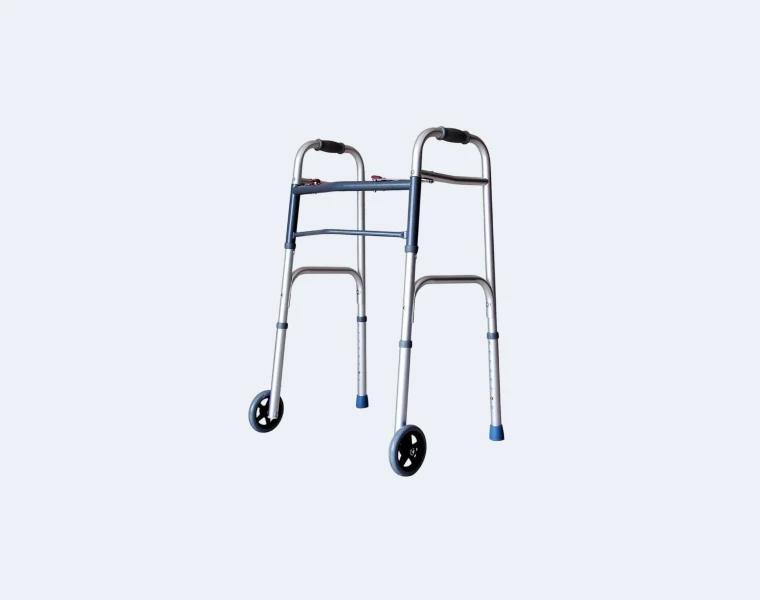
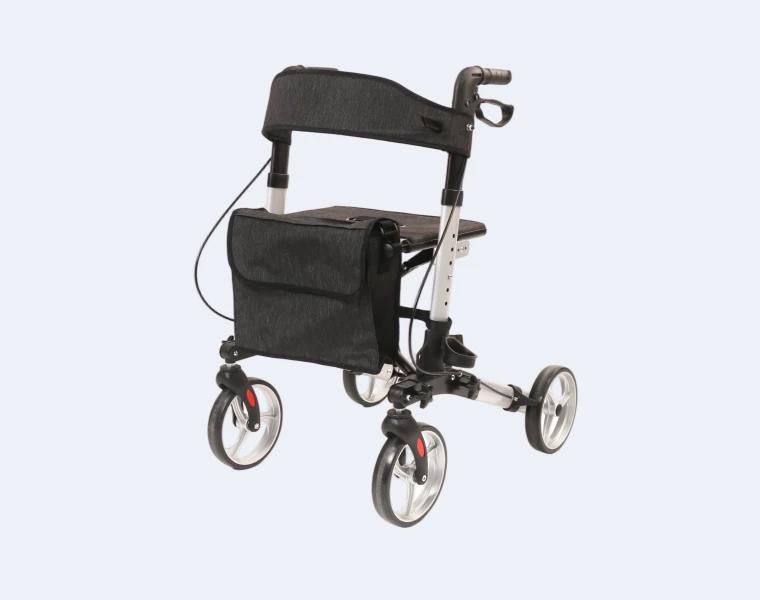
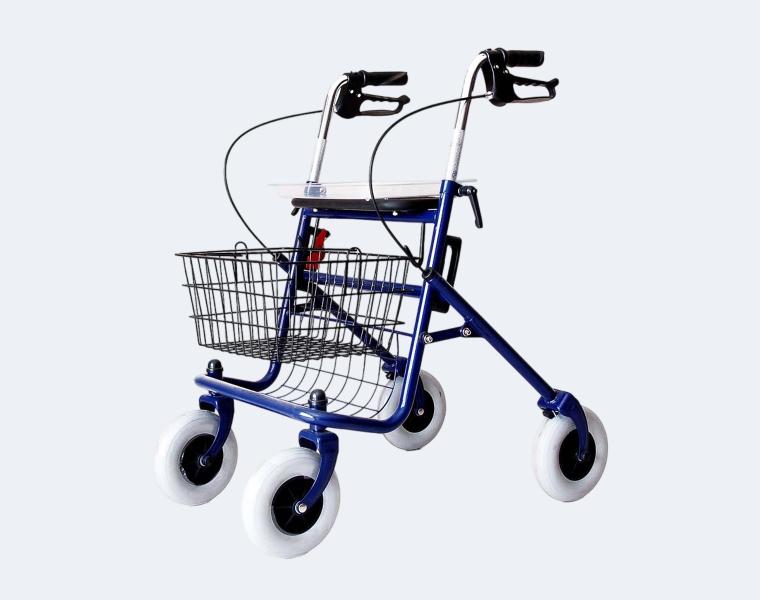
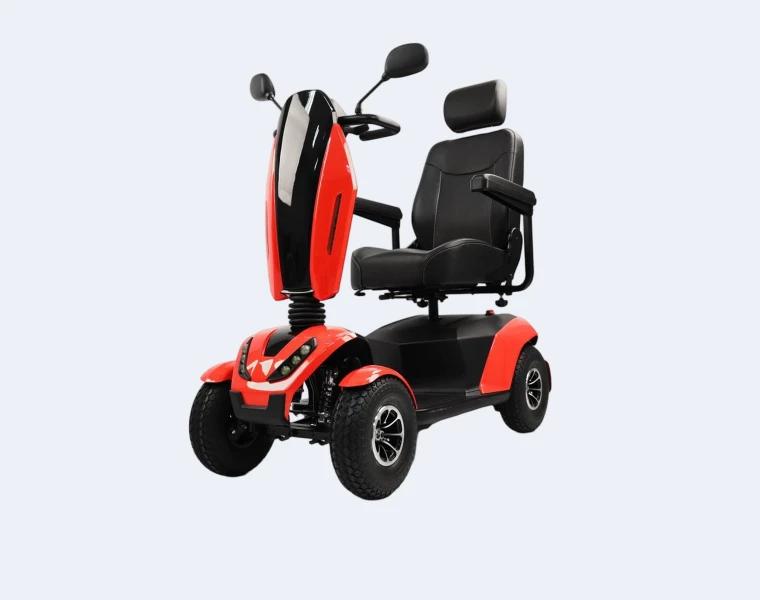
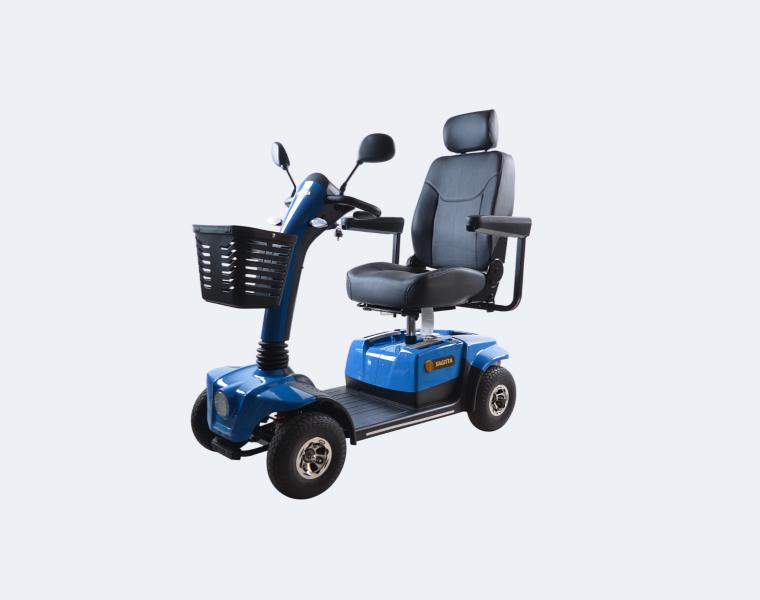
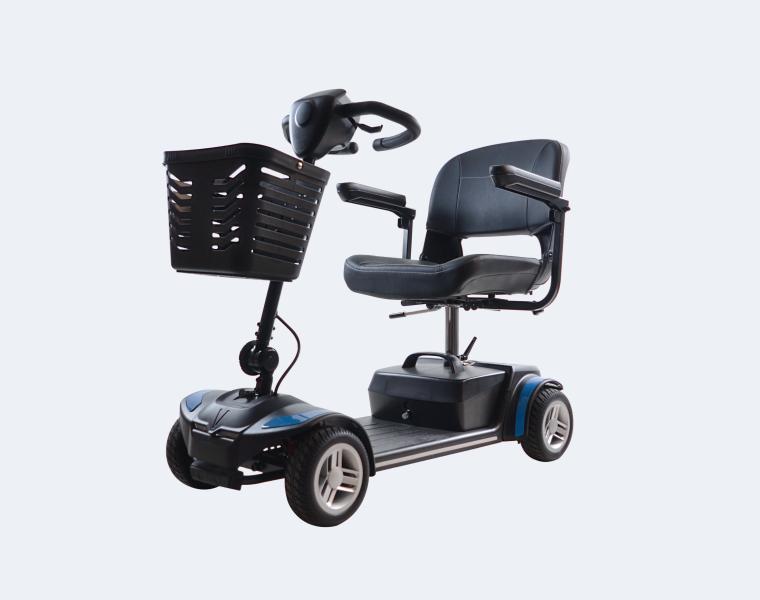
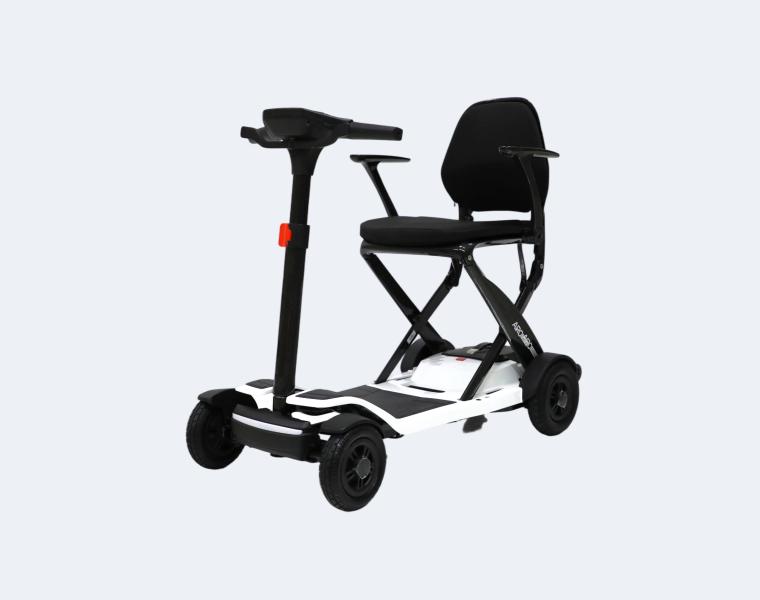
INTCO Medical’s rehabilitation equipment portfolio includes manual wheelchairs, powered wheelchairs, walkers, crutches, and other mobility support products. The applicable U.S. regulatory requirements vary by device type and classification.
Selected powered wheelchair products have associated FDA 510(k) records. For example, FDA 510(k) record K202482 is associated with the Y207 powered wheelchair under product code ITI. Additional FDA 510(k) records have been identified for selected powered wheelchair products.
Manual wheelchairs and certain walking aids may follow different regulatory pathways, including applicable 510(k) exemptions. FDA establishment registration and device listing records may also apply, depending on the device and responsible establishment.
Buyers should verify the exact model, legal manufacturer, device classification, applicable clearance or exemption status, product code, and current U.S. labeling. A clearance associated with one wheelchair model does not automatically apply to other models in the same product family.
Other Medical Consumables and Devices
In addition to examination gloves and mobility equipment, INTCO Medical supplies other medical consumables and devices, including medical face masks, protective apparel, ECG electrodes, and related products.
Selected medical surgical mask products have associated FDA 510(k) records. For example, FDA 510(k) record K202708 relates to a medical face mask under product code FXX.
Certain medical isolation gowns and related protective apparel may be classified as devices exempt from 510(k) requirements, subject to applicable regulations and exemption limitations. Registration and listing records should be verified where required.
For ECG electrodes, historical FDA 510(k) records and device listing information have been identified. However, the applicable regulatory status must be confirmed for each specific electrode model and intended use.
These examples illustrate the importance of product-specific regulatory verification. Buyers should not assume that FDA registration, device listing, or a historical clearance establishes clearance or approval for every product currently offered by a manufacturer.
Cold and Hot Therapy Products
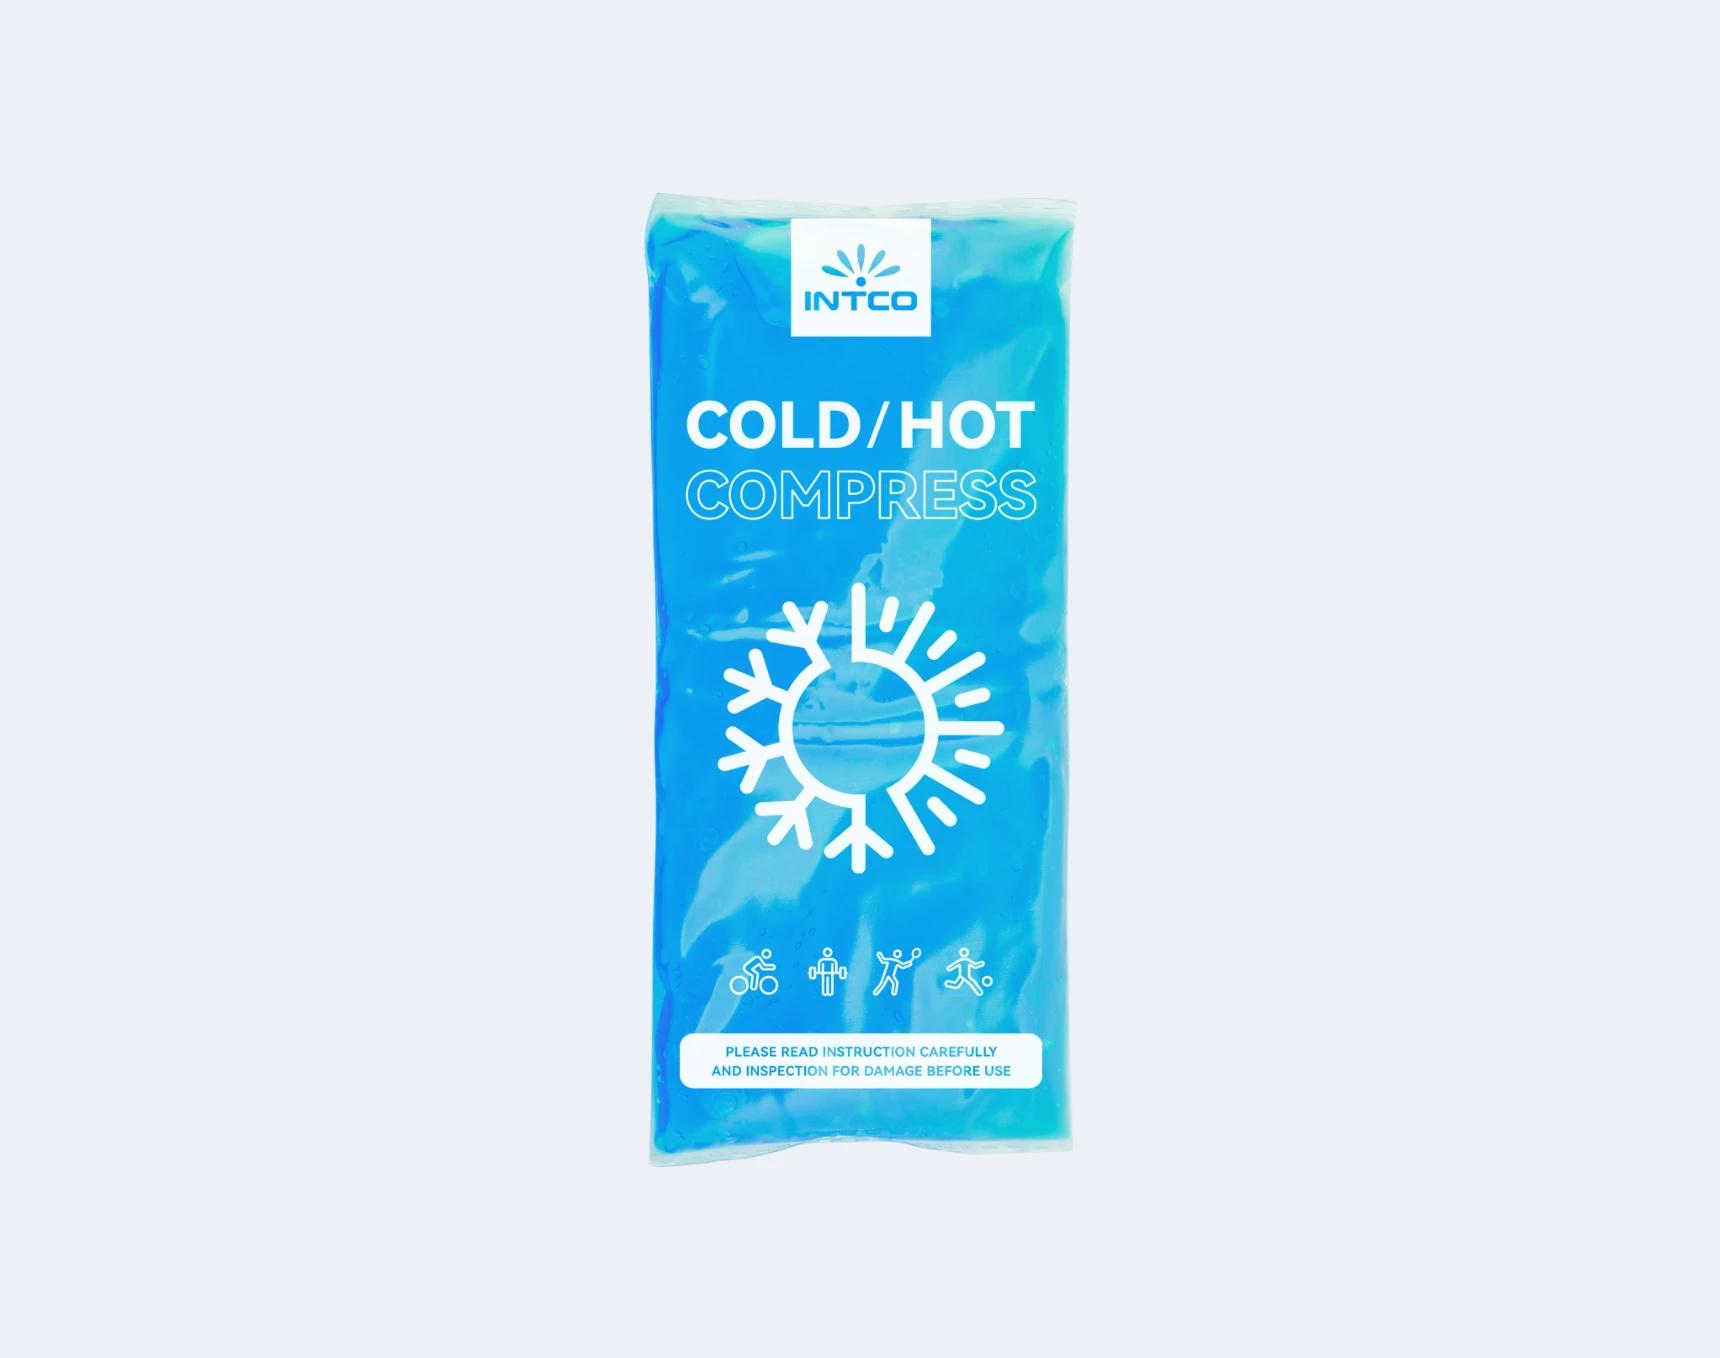
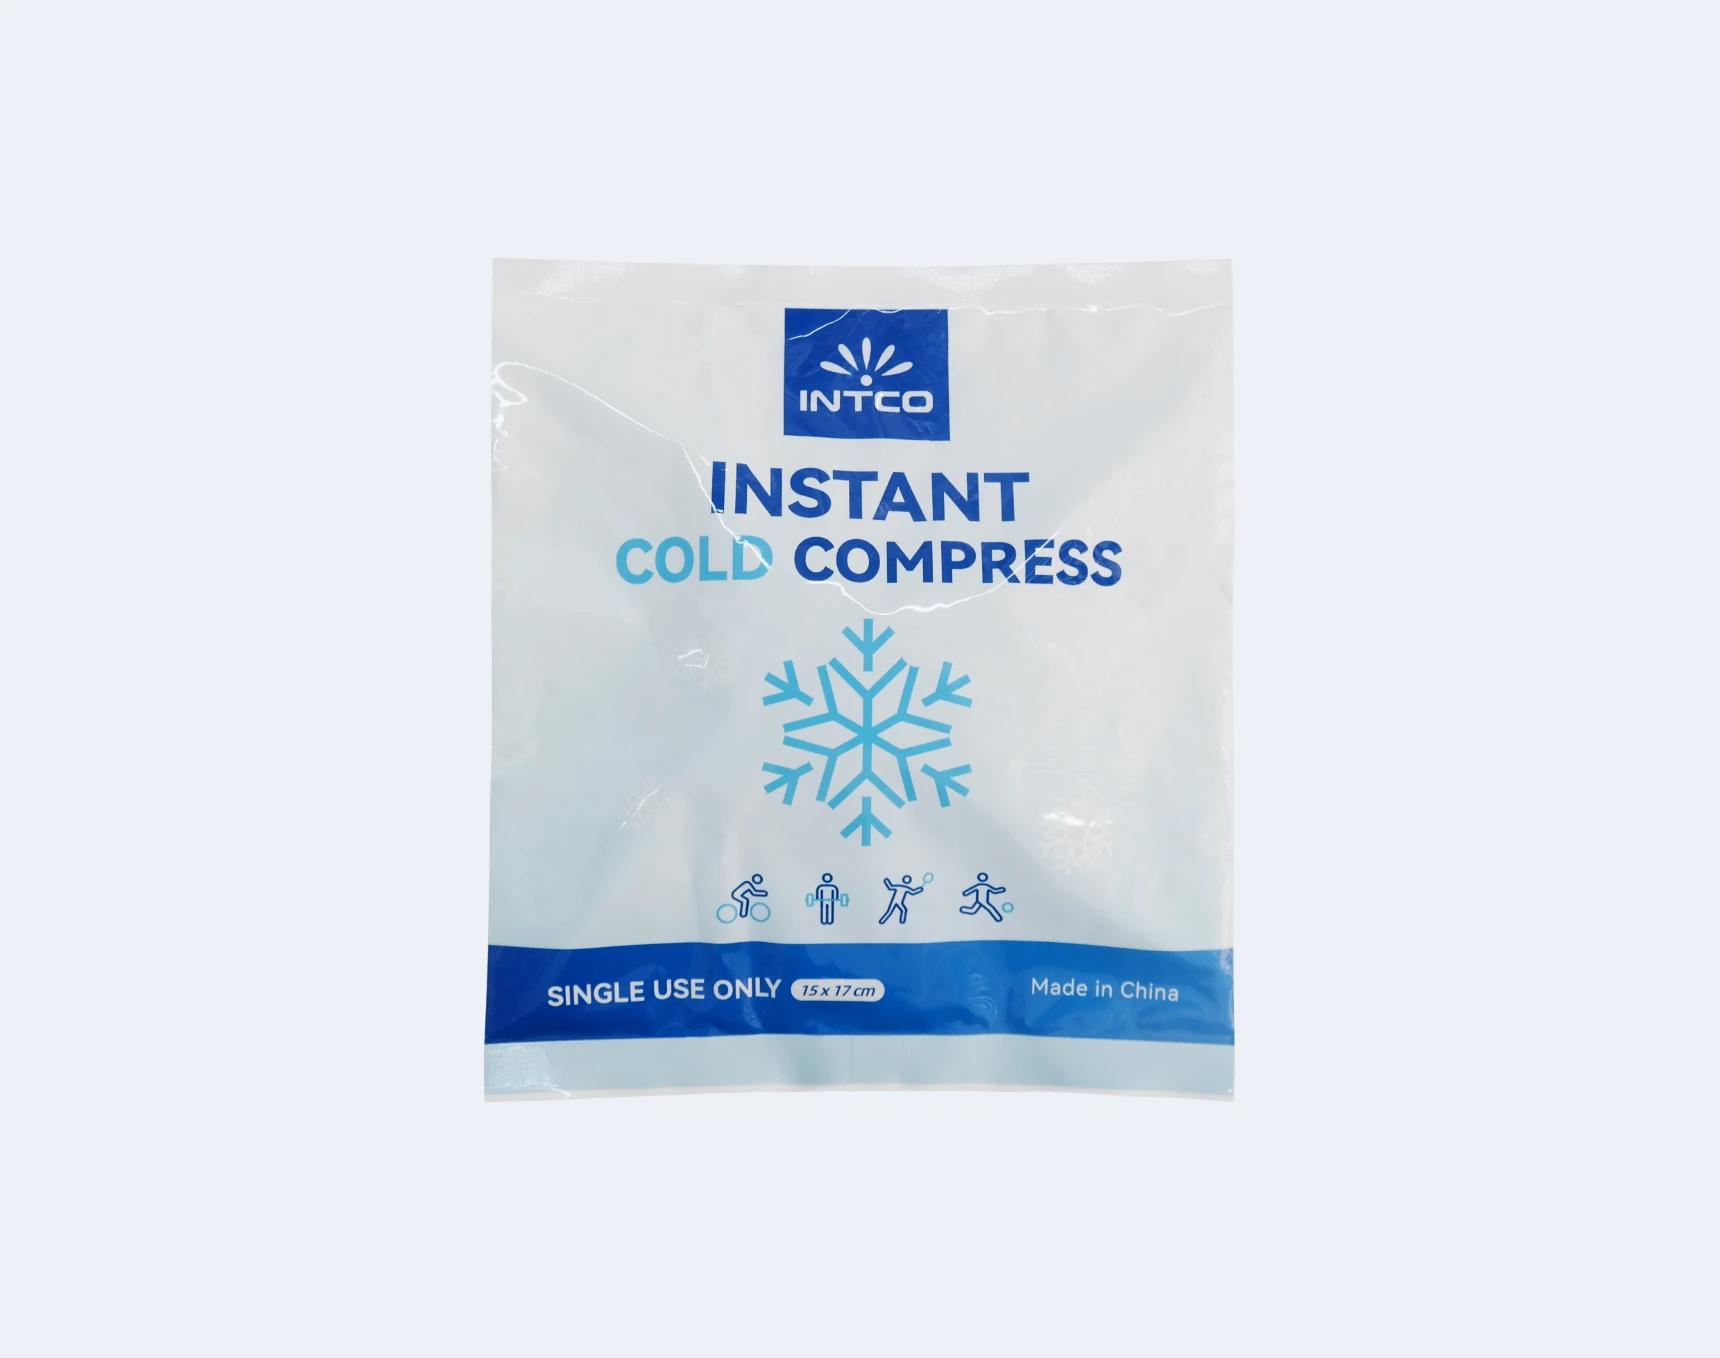
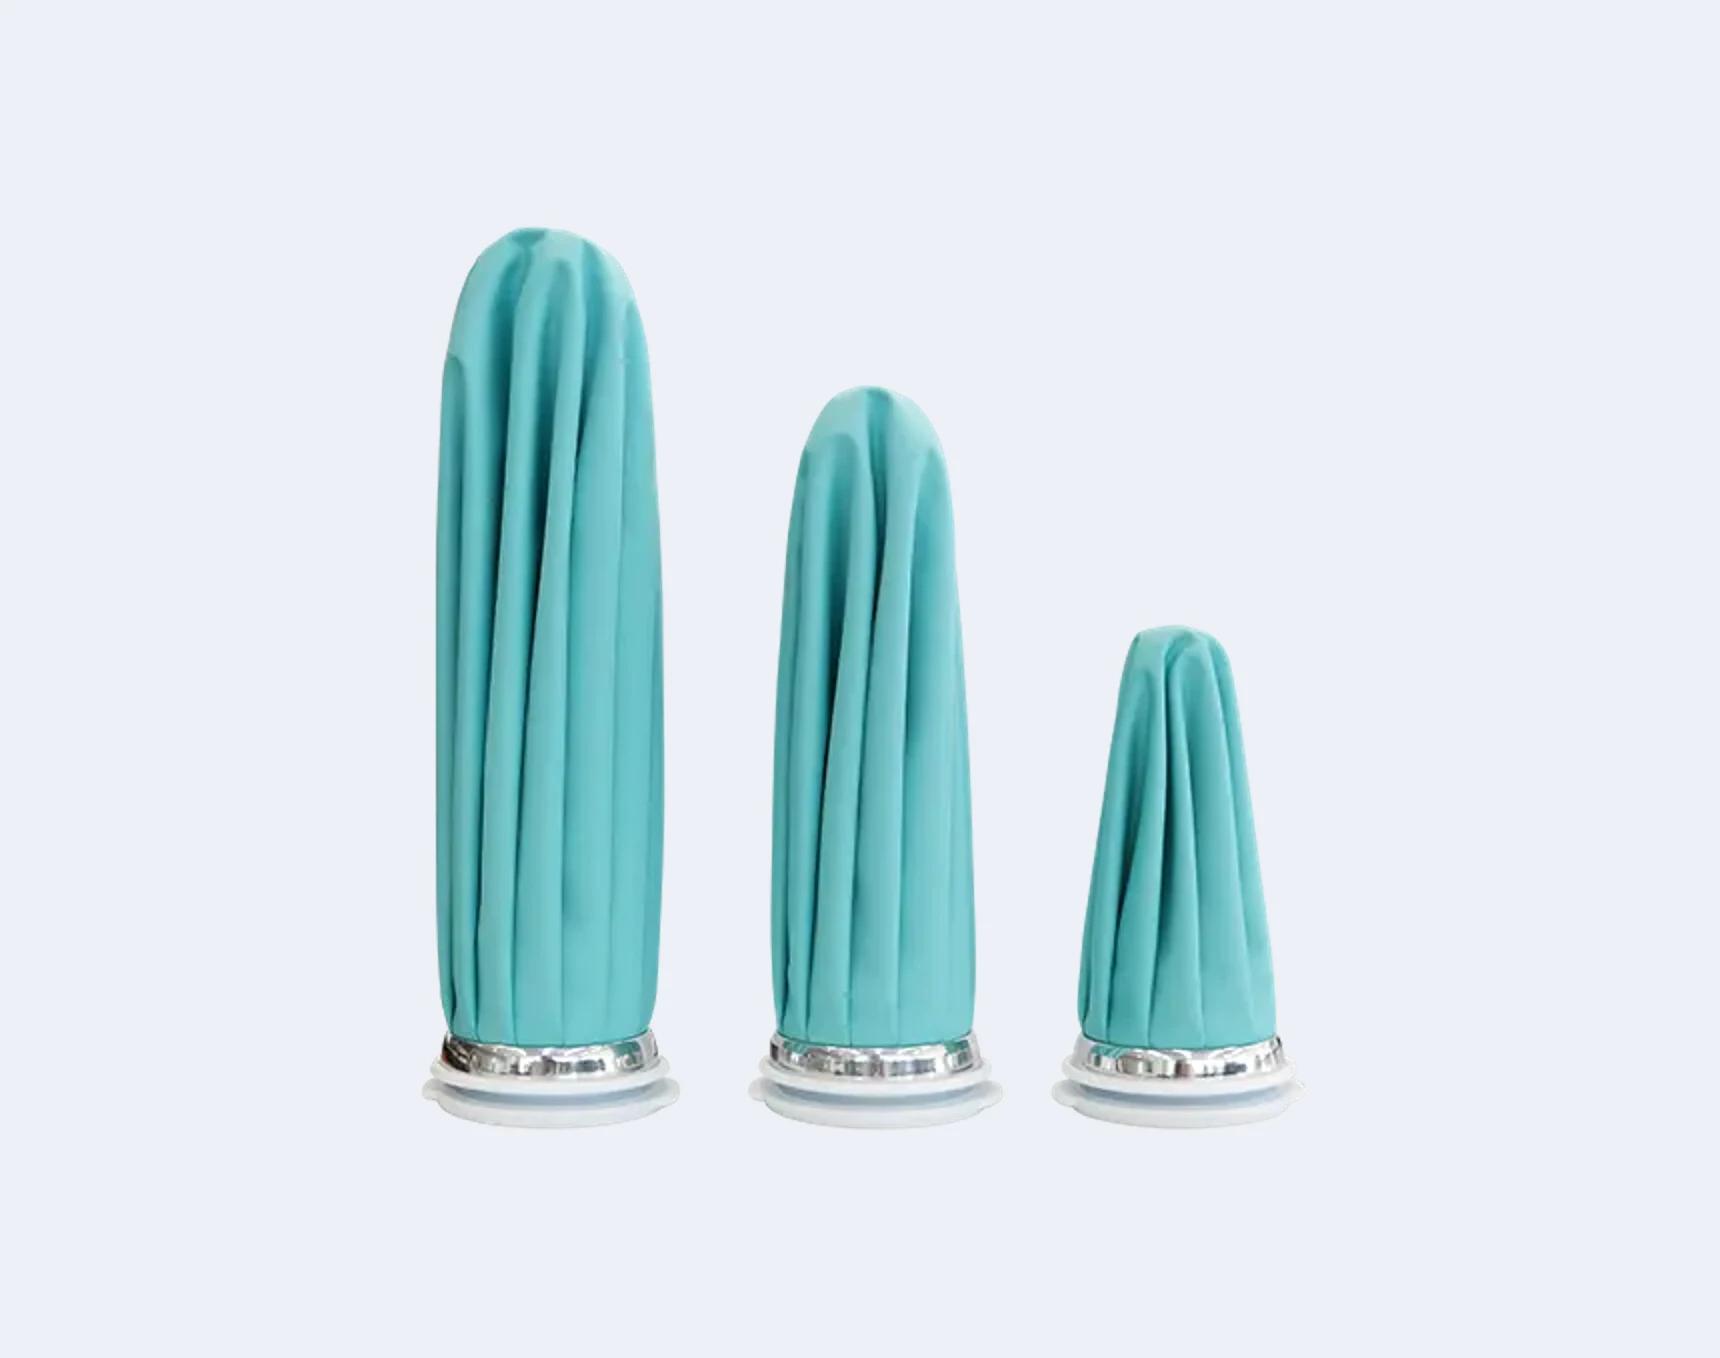
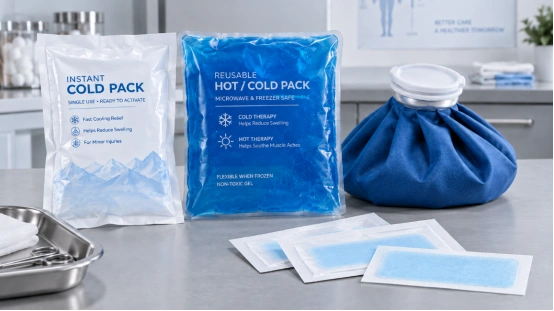
INTCO Medical also provides cold and hot therapy products, including instant cold packs, instant hot packs, reusable gel packs, ice bags, and fever cooling patches. These products may follow different FDA regulatory pathways depending on their classification and intended use.
FDA establishment registration and device listing information has been identified for selected cold and hot therapy products. The applicability of any 510(k) exemption should be verified for each specific product. The FDA status of fever cooling patches has not been independently confirmed.
The following catalogue information provides a starting point for further product-level regulatory review.
Product family | Catalogue details | Buyer follow-up |
Instant cold pack | 5 x 6 in and 6 x 9 in, disposable, external cold therapy, activated without refrigeration | Confirm FDA establishment registration and device listing, applicable classification and 510(k) exemption status, current labeling, and supporting test documentation |
Ice bag | 5, 6, 9, and 11 in, reusable ice bag with a fabric cover and secure closure | Confirm FDA registration and listing, applicable 510(k) exemption status, closure integrity, cleaning instructions, and current labeling |
Instant hot pack | 5 x 6 in and 6 x 9 in, disposable, activated without external heating | Confirm the applicable FDA registration and listing records, device classification, 510(k) exemption status, intended use, warnings, and test documentation |
Reusable gel pack | Reusable hot and cold therapy pack, available with PE/nylon or fabric materials, for applicable home-care use | Confirm FDA registration and listing, 510(k) exemption status, material specifications, leakage resistance, thermal performance, and cleaning instructions |
Fever cooling patch | 5 x 12 cm and 9.5 x 13 cm, disposable cooling patch; the catalogue describes a cooling effect lasting up to 8 hours | Confirm the applicable U.S. regulatory classification and FDA status. Request supporting evidence for cooling duration, skin-contact safety, stability, and labeling claims. |
Product Portfolio and Regulatory Scope
INTCO Medical’s portfolio spans disposable medical consumables, protective products, rehabilitation equipment, medical electrodes, and cold and hot therapy products. These categories are subject to different U.S. regulatory requirements, and product-specific evidence is essential for supplier qualification.
For buyers sourcing across multiple categories, the key consideration is not a manufacturer’s general FDA-related claims, but whether each product has the appropriate regulatory documentation for its intended use and U.S. distribution pathway.
How Does INTCO Medical Support a Multi-SKU U.S. Program?
INTCO Medical is a publicly listed medical products manufacturer with research and development, manufacturing, and global marketing operations across medical consumables and rehabilitation equipment. The company serves customers in more than 150 countries and regions and has over 15,000 customers worldwide. Its manufacturing and global supply capabilities support international procurement programs. However, supplier qualification for the U.S. market must still be based on the applicable regulatory documentation and quality evidence for each product.
Supplier Structure and Document Control
When you assess a medical device manufacturer supplying the U.S. market, review how its product, quality, regulatory, and logistics teams coordinate documentation and compliance. You need one controlled specification for each SKU, a current technical file, approved artwork, and lot traceability that connects production records to the shipment. A broad portfolio can support a coordinated program when each device keeps its own file.
When evaluating INTCO Medical products for the U.S. market, buyers should first confirm the applicable FDA regulatory pathway for each device. Ask which legal entity is responsible for the relevant regulatory records, which facility manufactures the product, and which parties handle applicable U.S. regulatory and postmarket responsibilities. Keep the verified information in the supplier qualification record.
Which Documents Should a 510(k) Medical Device Supplier Provide?
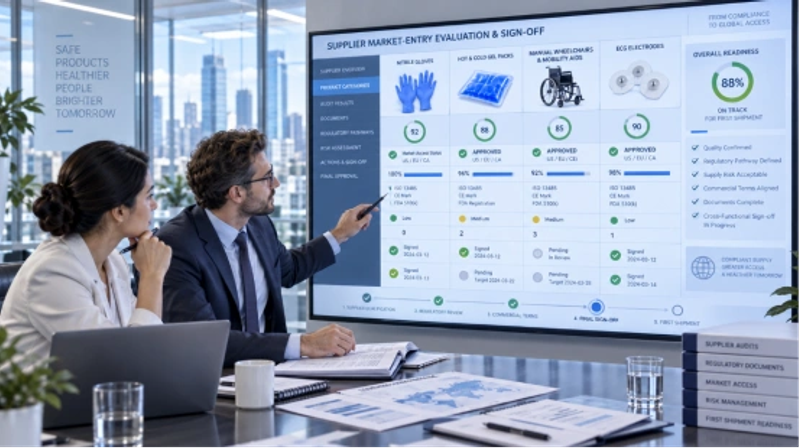
Your request should follow the product’s intended use and U.S. distribution role. A medical device supplier should be able to identify the applicable FDA regulatory pathway, provide supporting documentation for its product claims, and demonstrate that the commercial product matches the controlled product specifications and current labeling.
Core Request List
Document or data point | Why it matters to you |
Exact SKU, model, and commercial label | Links the purchase decision to one device configuration |
K number and FDA clearance decision, or applicable 510(k) exemption documentation | Verifies the applicable premarket requirements, clearance status, or exemption basis |
Registration and listing details | Identifies the establishment and listed device |
FDA product code and predicate device information, where applicable | Supports regulatory classification and substantial-equivalence review where applicable |
Current U.S. labeling and IFU | Checks claims, warnings, symbols, and use instructions |
ISO 13485 certificate scope | Shows which site and activities sit inside the quality system |
Technical file, test reports, and material declarations | Supports quality review and customer acceptance |
Lot traceability and release records | Connects delivered cartons to production and inspection data |
If you act as a U.S. medical device importer, clarify the responsibilities of the foreign manufacturer’s U.S. agent, the initial importer, and the importer of record, as applicable. Establish clear procedures for complaint handling and relevant change notifications. FDA establishment registration, device listing, and 510(k) clearance serve different regulatory purposes and should be reviewed separately.
How Should You Verify an INTCO SKU Before Purchase?
The review works best when you move from the product page to the record, then back to the physical pack. This sequence catches small mismatches, such as a size listed on a quote that does not appear on the current label.
Five Practical Checks
- Identify the intended user, care setting, intended use, target channel, and claimed benefit.
- Match the model, size, material, pack count, and label to the catalogue page and quotation.
- Check FDA establishment registrationand device listing records, confirm the device classification and applicable 510(k) requirements, and request the clearance number or exemption documentation where relevant.Compare the record with the technical file, labeling, IFU, test reports, and packaging artwork.
- Compare the record with the technical file, labeling, IFU, test reports, and packaging artwork.
- Approve a production-lot sample and keep the decision, lot code, and release documents together.
Private-label work needs a second pass. Confirm which legal entity is responsible for the applicable FDA regulatory records, who is responsible for the labeling, and how changes to materials, manufacturing sites, packaging, or intended use are assessed and reported when required. Certain changes may affect the applicable regulatory documentation or submission requirements.
How Can INTCO Medical Fit a U.S. Device Supply Strategy?
The catalogue presents a supplier that connects disposable protection, cold and hot therapy, and rehabilitation equipment. This combination supports coordinated supplier qualification across related product categories, while each SKU requires its own applicable U.S. regulatory classification, intended use, labeling, test evidence, and release documentation. The INTCO Medical product portfolio lets you map those categories before you request samples and regulatory documents.
For a buyer, the useful next step is specific. Send a model list, target claims, market role, package format, and forecast. Request the current U.S. regulatory and product documentation for each product line. The INTCO Medical corporate site provides the formal company contact route for that request.
FAQ
Q1: Does the catalogue prove that every INTCO Medical product has FDA 510(k) clearance?
A: No. FDA-related marks in a product catalogue do not establish 510(k) clearance or FDA approval. Some products may be subject to FDA registration and listing and may qualify for a 510(k) exemption, while others may require a premarket submission. Buyers should verify the regulatory pathway and supporting records for each specific product.
Q2: What is the difference between FDA medical device registration and listing and 510(k) clearance?
A: FDA establishment registration and device listing identify registered establishments and listed devices but do not indicate FDA approval or clearance. A 510(k) clearance is a separate premarket decision based on substantial equivalence. Certain devices may be exempt from 510(k) requirements. Buyers should verify which requirements apply to the specific product.
Q3: Which INTCO Medical product categories have FDA-related regulatory records?
A: FDA-related regulatory records have been identified for selected INTCO Medical products across several categories, including medical examination gloves, powered wheelchairs, medical surgical masks, ECG electrodes, and cold and hot therapy products. Depending on the specific device, these records may involve 510(k) clearance, establishment registration and device listing, or applicable 510(k) exemptions. Buyers should verify the legal manufacturer, current model, product code, regulatory status, and supporting FDA documentation for each product.
Q4: What should a U.S. medical device importer request before ordering?
A: Request the exact SKU and current label, applicable FDA registration and listing information, device classification, 510(k) clearance or exemption evidence where relevant, ISO 13485 certificate scope, instructions for use, test reports, packaging artwork, and lot traceability documentation.
Q5: How does a medical consumables manufacturer support a multi-category program?
A: It can coordinate specifications, artwork, samples, quality records, and shipment planning across gloves, masks, therapy products, and related lines. Each SKU still needs its own classification, intended use, evidence, and release decision.
Q6: Does FDA 510(k) clearance for one INTCO Medical product cover other models or product categories?
A: No. FDA 510(k) clearance applies to the specific device and indications for use covered by the relevant clearance decision. It does not automatically extend to all models within a product family or to other product categories. Buyers should verify whether the exact product configuration is covered by the applicable clearance or follows another regulatory pathway.
Q7: How can U.S. buyers verify FDA records for INTCO Medical products?
A: Buyers can consult the FDA’s official 510(k), establishment registration, and device listing databases. The review should include the legal manufacturer, device name, product code, K number where applicable, intended use, and relevant model information. Buyers should also request current product-specific documentation from INTCO Medical before making purchasing or regulatory decisions.