INTCO in Europe: Compliance, Distribution, and Market Support
Medical Device Quality Control: What Buyers Should Verify Before Supplier Approval
FROM: INTCO Medical
|
02 Oct 2026
Supplier approval should rest on records that connect the quality system to the product you plan to buy. A certificate starts the review. Process data, test methods, lot history, and response times show whether medical device quality control works during production. Build an evidence trail from incoming material to released shipment and field complaint.
What Should You Verify in the Quality Management System?
A quality management system sets the rules for document control, production, inspection, complaints, and corrective action. During qualification, check how those rules operate on your device family and at the proposed manufacturing site.
Certificate Scope and Audit Evidence
For an ISO 13485 supplier, request the current certificate, issuing body, expiry date, covered sites, and scope. Match the scope to the product and activity in your purchase plan. Ask for the latest external-audit summary, open findings, closure evidence, and internal-audit schedule. This quality audit evidence belongs in the approval file.
For US-bound devices, confirm the site follows the FDA Quality Management System Regulation, effective February 2, 2026, and its applicable device requirements.
CAPA and Management Review
Review how the supplier manages corrective and preventive actions, including root-cause analysis, corrective action, and effectiveness verification. Confirm that management review covers relevant quality-system performance and follow-up actions. Detailed records may be reviewed where appropriate under applicable confidentiality arrangements. Recurring defects after closure may indicate that corrective actions or effectiveness checks require further review.
Which Production Controls Prove Repeatable Output?
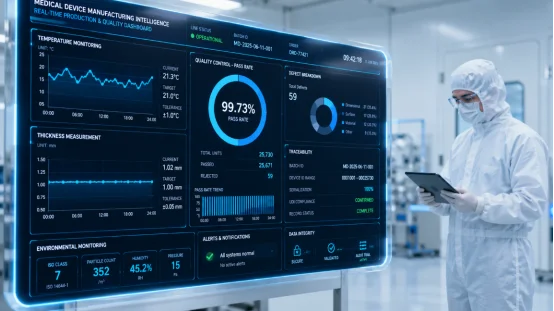
Factory equipment matters when approved settings, trained operators, maintained tools, and validated processes govern its use. Your medical device supplier audit should follow one representative lot through the actual production route. That walk often reveals more than a polished conference-room presentation.
Material, Process, and Release Gates
Map three linked control stages:
|
Control Gate |
Evidence to Request |
Buyer Check |
|
Incoming quality control |
Material specification, supplier status, sampling record, test result |
Lot identity matches the production record |
|
In-process quality control |
Approved parameters, first-piece check, defect log, equipment status |
Limits and reaction plans are recorded |
|
Final quality control |
Finished-product results, packaging check, release signature |
All acceptance criteria have results |
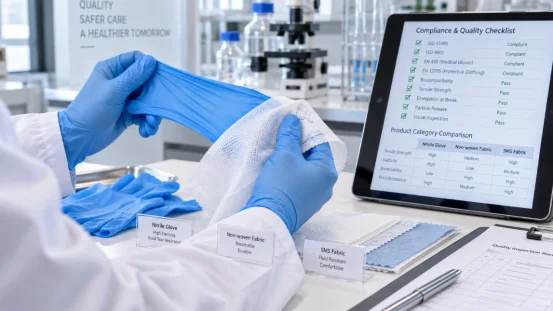
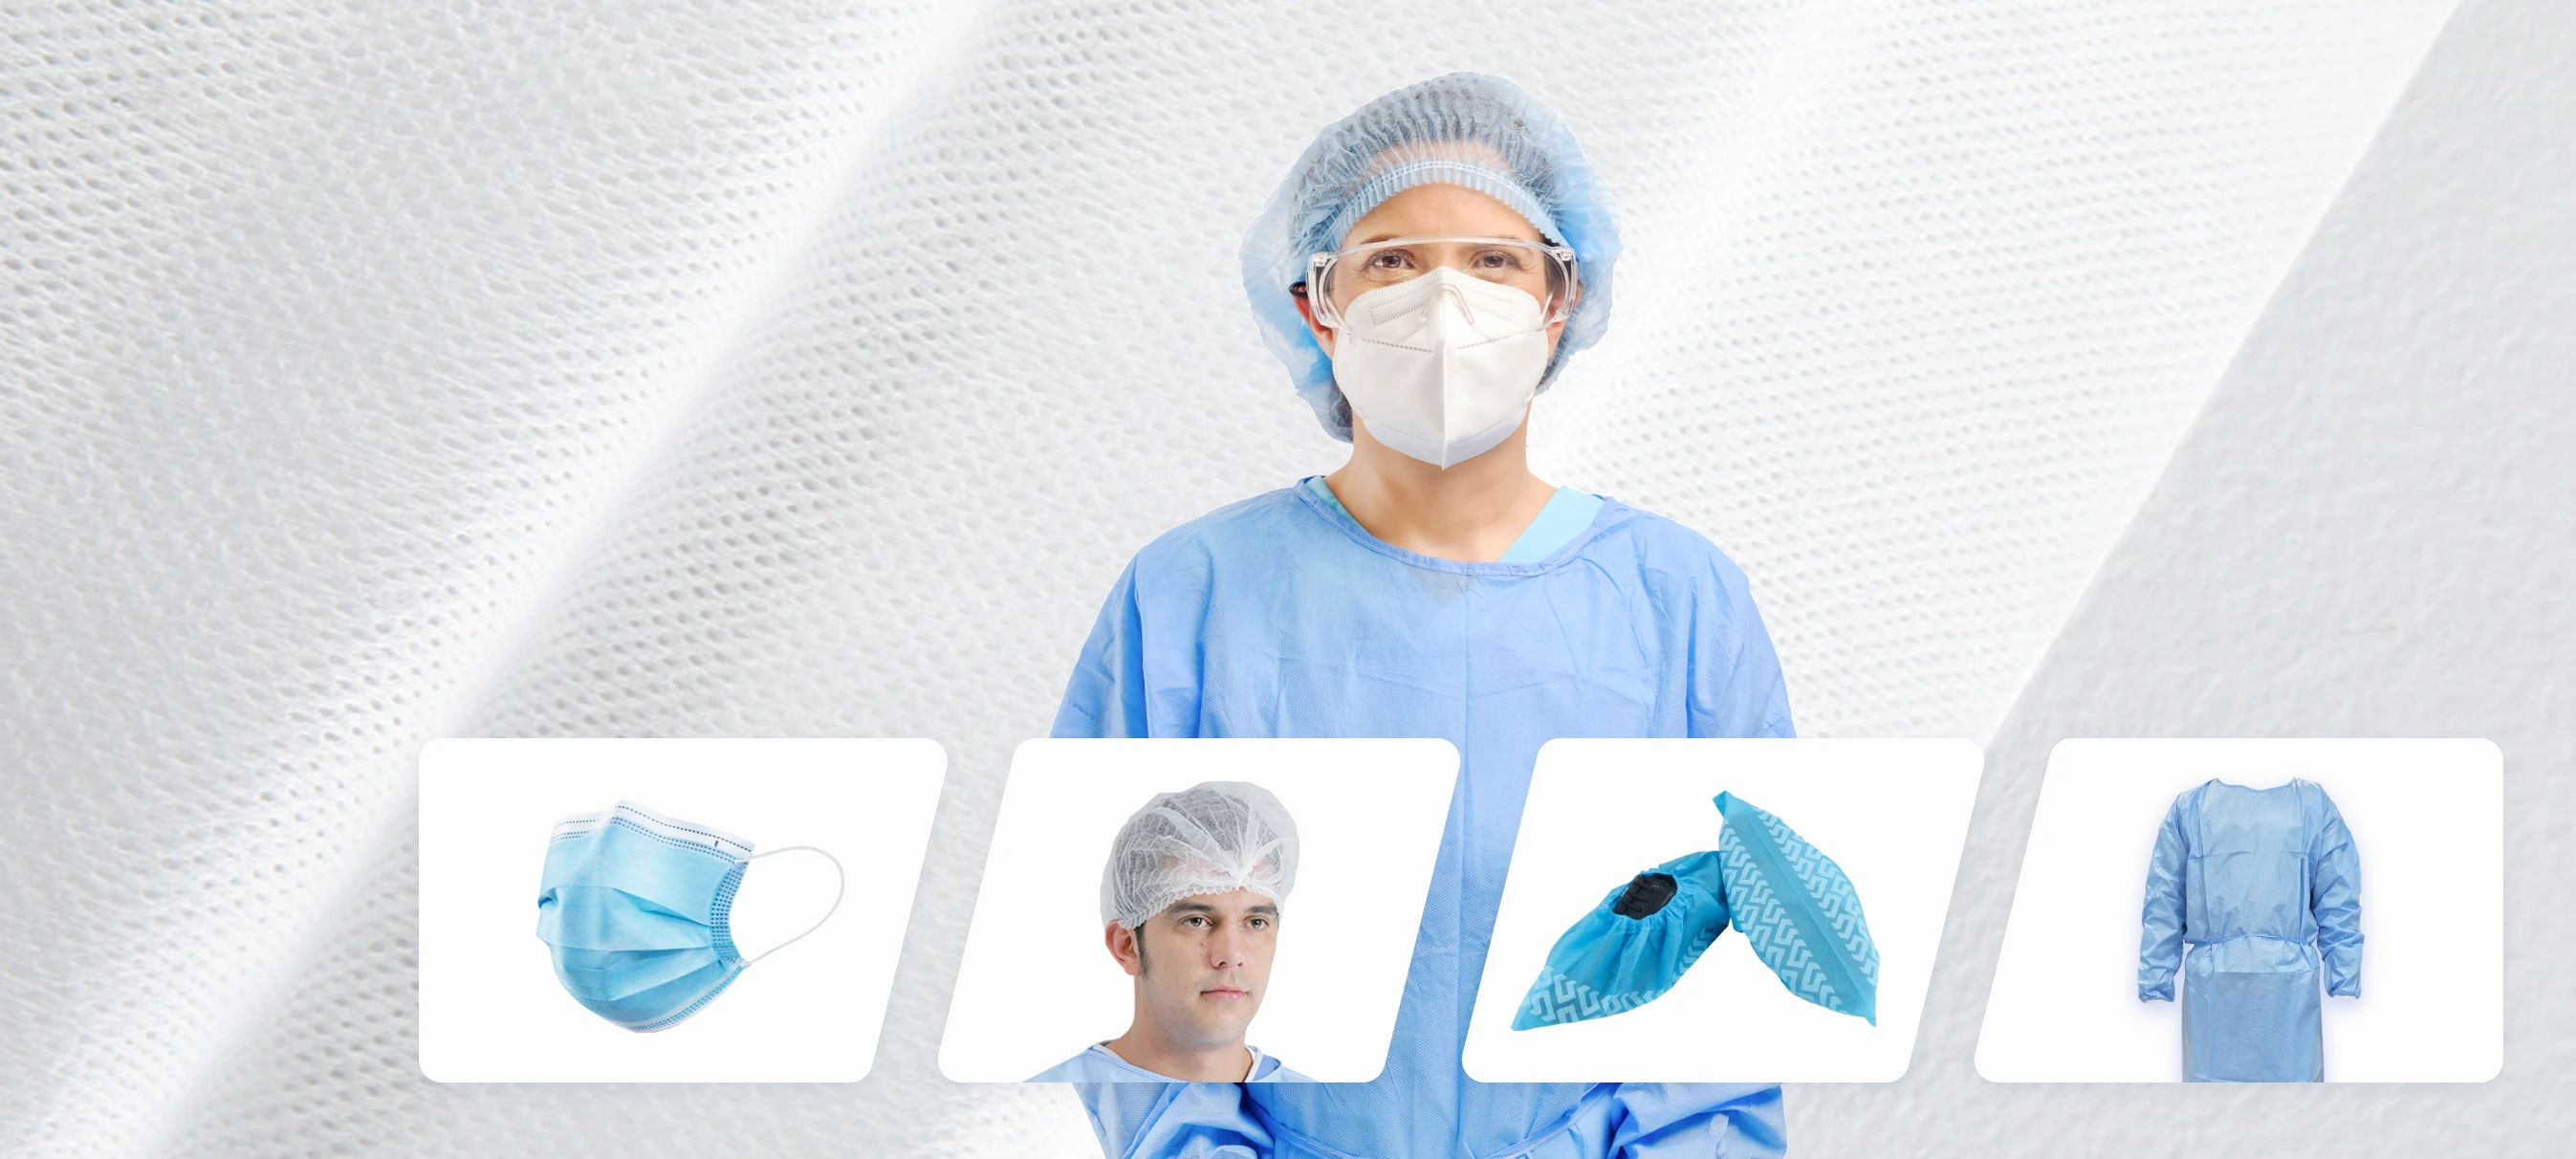
For a disposable glove lot, for example, connect raw-material identity with dimensions, visual inspection, tensile data, and leak-test results. For masks or protective apparel, use the stated intended use to select filtration, breathability, seam, barrier, or attachment tests. The specification should name the method, limit, sample size, and disposition rule.
Validation and Measurement Control
Confirm that critical processes are appropriately validated when their output cannot be fully verified later. Review relevant validation evidence, where appropriate, under agreed confidentiality arrangements. This process validation should define approved operating limits. Check calibration status, measurement range, accuracy, and out-of-tolerance handling.
Sterile, patient-contacting, or shelf-life claims may also call for sterilization validation, ISO 10993 biocompatibility evidence, packaging validation, transport testing, and aging data. This applies to product families such as face masks when the specification carries filtration, fluid-resistance, or shelf-life claims. The evidence set must match the claim and intended use.
How Do You Test Traceability and Change Control?
Traceability becomes useful when your team can retrieve a full record quickly. Run a desk exercise during the audit: select a finished lot and ask the supplier to trace materials, line, date, operators, inspections, deviations, and release approval.
Lot Records and Recall Readiness
Effective batch traceability links the shipped unit or carton to production, test history, and lot release records. Set a retrieval target in the supplier quality agreement, then time the exercise. Ask the supplier to trace one raw-material lot forward into every affected finished lot as well. This second direction defines the potential recall population.
Controlled Changes
List changes that require written notice and buyer approval: material source, formulation, design, tooling, process parameters, test method, packaging, label, software, sterilization route, and manufacturing site. Set notice periods and required evidence. A small carton or label edit can affect registration records and market release. Busy launch teams sometimes miss that link.
What Regulatory and Product Evidence Belongs in the Approval File?
Company registrations show organizational status. Device-specific regulatory documentation shows whether the proposed model, variant, and intended use have the required route into your target market. Review documents against the legal manufacturer, site, product code, classification, and destination country.
Product-Level Document Matrix
Build one matrix for every SKU:
- approved specification and signed sample record
- applicable registration, listing, declaration, or certificate
- risk-management output aligned with ISO 14971
- product test reports and laboratory status
- label, instructions for use, language, UDI, and shelf-life evidence
- complaint, vigilance, and field-action responsibilities
Check names, model numbers, standards, dates, and report samples across the set. A valid company certificate cannot establish compliance for every device. Your approval file needs the records that cover the purchased item.
How Should You Score the Supplier Approval Decision?
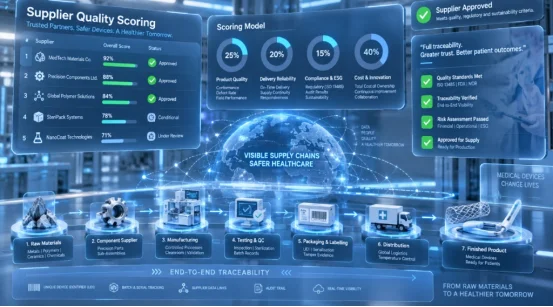
A weighted score keeps commercial pressure from quietly lowering quality thresholds. Set mandatory gates first, then score the remaining evidence. Supplier approval should require closure of critical gaps before the first commercial lot.
A Practical Approval Model
|
Review Area |
Suggested Weight |
Approval Evidence |
|
Quality system and audit history |
25% |
Current scope, audit status, corrective-action process |
|
Process and testing controls |
25% |
Control plan, validation, lot results |
|
Regulatory fit |
20% |
Product and market document matrix |
|
Traceability and change control |
15% |
Timed trace, notification procedure |
|
Complaint and supply response |
15% |
Escalation path, complaint-handling process, continuity plan |
Use a supplier qualification checklist with three outcomes: approved, conditionally approved with dated actions, or rejected. Record the rationale, owners, due dates, and re-evaluation interval. Add qualified sources to the approved supplier list with their permitted product and site scope. Price, lead time, and capacity belong in the sourcing decision, while critical quality or regulatory gaps remain approval gates.
Where Can Buyers Apply These Checks across Related Product Categories?
Once the evidence requirements are fixed, category coverage becomes useful because the same approval logic can support several disposable product lines. The supplier still needs a separate specification, risk file, test set, and release record for each SKU.
Connecting Category Breadth with Quality Evidence
INTCO Medical presents a product portfolio spanning disposable gloves, masks, protective apparel, and care products. That category range can help a buyer coordinate artwork, shipment planning, and document review through one program. Product acceptance stays category specific.
Define material, dimensions, physical tests, and leak criteria for gloves, then use separate filtration, construction, or barrier checks for masks and apparel. Ask for production-lot samples, controlled specifications, test records, and market documents tied to the exact items under review. This gives your approval team one working channel while preserving product-level evidence.
FAQ
Q1: What Is the First Document to Check during Supplier Approval?
A: Start with the ISO 13485 certificate, then verify its site, scope, issuer, and expiry against the proposed product and manufacturing activity.
Q2: What Should Buyers Check in a Supplier’s CAPA Process?
A: Review how complaints and nonconformities are investigated, how root causes are identified, and how corrective actions are verified for effectiveness. Detailed CAPA records may be subject to confidentiality requirements.
Q3: What Makes Batch Traceability Effective?
A: The system links a finished lot to materials, production conditions, inspections, deviations, release, and shipment records, and traces affected material forward.
Q4: Should a Certificate Replace a Factory Audit?
A: The certificate confirms a defined management-system scope. The audit tests how the site applies those controls to your product, process, and records.
Q5: When Should a Supplier Receive Conditional Approval?
A: Use it for limited, dated gaps with defined owners and controls. Critical regulatory, safety, traceability, or validation gaps require closure before commercial production.